Scalability
From a single review to global submission support, we scale effortlessly to meet your project needs.
Get FDA-ready Investigational New Drug applications delivered by regulatory experts trusted by biotech and pharma leaders. Complete pre-IND strategy, CMC documentation, nonclinical study summaries, and clinical protocol packages that accelerate your path to first-in-human studies. Not sure you're ready to file? Download our IND Readiness Checklist.
Criterion Edge provides end-to-end IND writing and regulatory support across the full IND lifecycle—from Pre-IND briefing packages to integrated nonclinical and clinical summaries, Investigator’s Brochure development, and CMC regulatory documentation. We partner with sponsors to translate complex development data into clear, FDA-aligned narratives that support first-in-human study authorization and confident regulatory decision-making.
We combine regulatory strategy, scientific rigor, and execution speed to deliver IND submissions that are clear, compliant, and built for FDA review.
We align early on product profile, indication, regulatory pathway, and FDA expectations—defining scope, timelines, and document strategy to support a successful IND.
We assess available nonclinical, clinical, and CMC data to confirm readiness, identify gaps, and define the structure and level of detail required for each IND section.
We develop clear, FDA-aligned narratives across nonclinical, clinical, and CMC sections, integrating pharmacology, toxicology, PK/TK, and manufacturing data into a cohesive regulatory story.
Each IND undergoes structured quality and consistency reviews to ensure alignment with FDA guidance, internal data sources, and eCTD requirements.
We prepare the IND for eCTD submission and support FDA interactions during the 30-day review period, including responses to information requests and clarification questions.
At Criterion Edge, we provide strategic, FDA-aligned IND writing support that helps sponsors move efficiently from preclinical development to first-in-human studies. Our approach combines regulatory insight, scientific rigor, and clear communication to support confident FDA engagement and clinical program initiation.
Clients are supported by experienced regulatory medical writers who lead IND strategy, content development, and coordination across nonclinical, clinical, and CMC sections—ensuring a focused, defensible submission tailored to each program
We develop and integrate nonclinical, clinical, and CMC content within the IND and eCTD structure, ensuring consistency across study narratives, dose justification, safety rationale, and manufacturing descriptions, aligned with FDA guidance and current review expectations.
Our team synthesizes pharmacology, toxicology, PK/TK, clinical, and CMC data into clear, submission-ready narratives, supported by structured quality and compliance reviews to minimize FDA questions and avoid delays to clinical start.
We translate complex scientific data into persuasive regulatory narratives that support first-in-human dosing, enable FDA review, and help sponsors advance their clinical programs with confidence.
From initial IND filings to amendments and safety updates, Criterion Edge delivers comprehensive IND documentation that meets FDA requirements and accelerates your path to clinical trials.
Quality Assurance Manager
PhD's, MD's, Scientists, Researchers
Medical Librarian, Clinical Evidence Experts
Principal Writer / Strategic Leadership
Project Management
Your literature review deserves more than just a freelancer. At Criterion Edge, each project is staffed with a dedicated, specialized team to ensure speed, consistency, and subject-matter accuracy.

We tap into a distributed team with deep therapeutic and regulatory experience - working around the clock to move your project forward.
You'll work with protocol developers, medical writers, and therapeutic-area experts aligned to your specific evidence needs.
Need to move faster or adjust scope? We scale resources up or down to meet your timeline and budget - without sacrificing quality.
We don't just check boxes - we partner with your team to deliver high-quality, submission-ready results. Here's what sets us apart:
From a single review to global submission support, we scale effortlessly to meet your project needs.
Our team includes seasoned experts across regulatory, clinical, and therapeutic domains.
Built-in quality control ensures every deliverable is compliant, consistent, and audit-ready.
Streamlined workflows and dedicated project teams reduce turnaround time - without cutting corners.
Below are the average scores (out of 5) for each of the criteria we ask our clients to grade us on after every project.

Pharmaceutical, in-vitro diagnstics and medical device companies face growing pressure to deliver high-quality, compliant submissions on tight timelines - with limited internal resources. Criterion Edge was built to fill that gap.
Navigating evolving requirements like EU MDR and FDA guidance takes expertise and constant vigilance. We simplify the process for you.
Late reports, misaligned expectations, or unclear deliverables waste valuable time. Our team is known for quality, clarity, and on-time delivery.
Internal writing teams can be costly and hard to scale. Outsourcing with us helps you meet your evidence goals while staying within budget.
Medical writing and regulatory expertise is niche and hard to hire. With decades of experience, our consultants, writers and reviewers are ready to support your projects right now.
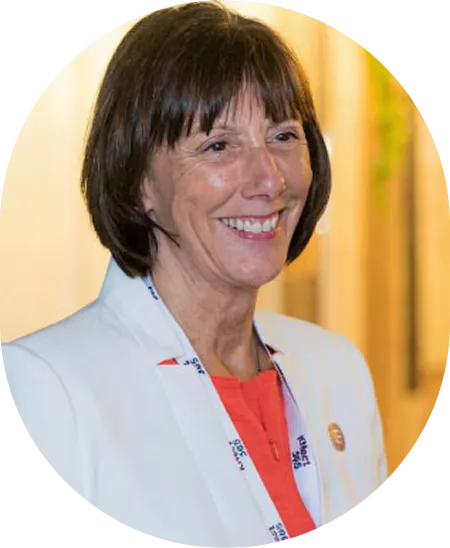
With over two decades in the pharmaceutical and medical device sectors, Laurie Mitchell founded Criterion Edge in 2016 with the goal to build a world-class team of industry experts dedicated to providing regulatory support and comprehensive writing services to pharma, biotech and device clients worldwide.
Under her leadership, Criterion Edge has supported top global brands across regulatory submissions, clinical strategy, and market access. Today, Criterion Edge has grown into a recognized leader, trusted by clients ranging from ambitious start-ups to some of the largest companies in the world, but our dedication to quality, scientific rigor and regulatory excellence has not changed.
Let’s discuss your Investigational New Drug Application needs and how Criterion Edge can support your strategic objectives.