Scalability
From a single review to global submission support, we scale effortlessly to meet your project needs.
Get FDA-ready 510(k) packages delivered by regulatory experts trusted by leading medtech companies. Comprehensive predicate analysis, substantial equivalent documentation, and performance testing summaries that reviewers expect. Not sure you're ready to submit? Download our 510(k) Readiness Checklist.
Criterion Edge offers specialized services across the entire 510(k) submission lifecycle—from substantial equivalence documentation and predicate analysis to performance testing summaries, and strategic consulting for medical device clearance.
We deliver efficient, FDA-ready 510(k) submissions through a transparent, collaborative process designed to reduce risk and keep programs moving.
We align early on device classification, regulatory pathway, predicate strategy, and submission goals to set a clear, defensible path forward.
We review your existing device documentation, testing data, and supporting evidence to confirm readiness and identify any gaps before drafting begins.
We integrate technical and clinical evidence and build clear, persuasive, substantial equivalence arguments aligned with FDA expectations.
We develop a cohesive 510(k) within FDA’s eSTAR template, applying rigorous quality and compliance reviews to ensure clarity, consistency, and reviewability.
We finalize and support submission to the FDA and remain engaged through review, including management of FDA questions and Additional Information requests.
At Criterion Edge, we provide strategic, FDA-aligned 510(k) support that helps medical device manufacturers move efficiently from development to clearance. Our approach combines regulatory insight, technical rigor, and clear communication to support confident regulatory and market entry decisions.
Clients are supported by experienced regulatory writers who lead predicate identification, substantial equivalence strategy, and submission development—ensuring a focused, defensible approach tailored to each device.
We develop and integrate technical, nonclinical, and clinical content within FDA’s eSTAR framework, ensuring consistency across device description, performance data, and substantial equivalence arguments, aligned with applicable FDA guidance and recognized standards.
Our team synthesizes device data, testing results, and supporting documentation into clear, submission-ready content, supported by structured quality and compliance reviews to minimize review risk and FDA follow-up.
We translate complex technical information into persuasive regulatory narratives that support substantial equivalence, streamline FDA review, and help manufacturers advance their products to market with confidence.

From Traditional to Special and Abbreviated pathways, Criterion Edge delivers FDA-aligned 510(k) submissions that demonstrate substantial equivalence and support efficient device clearance.
Quality Assurance Manager
PhD's, MD's, Scientists, Researchers
Medical Librarian, Clinical Evidence Experts
Principal Writer / Strategic Leadership
Project Management
Your 510(k) deserves more than just a freelancer. At Criterion Edge, each project is staffed with a dedicated, specialized team to ensure speed, consistency, and subject-matter accuracy.

We tap into a distributed team with deep therapeutic and regulatory experience - working around the clock to move your project forward.
You'll work with protocol developers, medical writers, and therapeutic-area experts aligned to your specific evidence needs.
Need to move faster or adjust scope? We scale resources up or down to meet your timeline and budget - without sacrificing quality.
We don't just check boxes - we partner with your team to deliver high-quality, submission-ready results. Here's what sets us apart:
From a single review to global submission support, we scale effortlessly to meet your project needs.
Our team includes seasoned experts across regulatory, clinical, and therapeutic domains.
Built-in quality control ensures every deliverable is compliant, consistent, and audit-ready.
Streamlined workflows and dedicated project teams reduce turnaround time - without cutting corners.
Below are the average scores (out of 5) for each of the criteria we ask our clients to grade us on after every project.

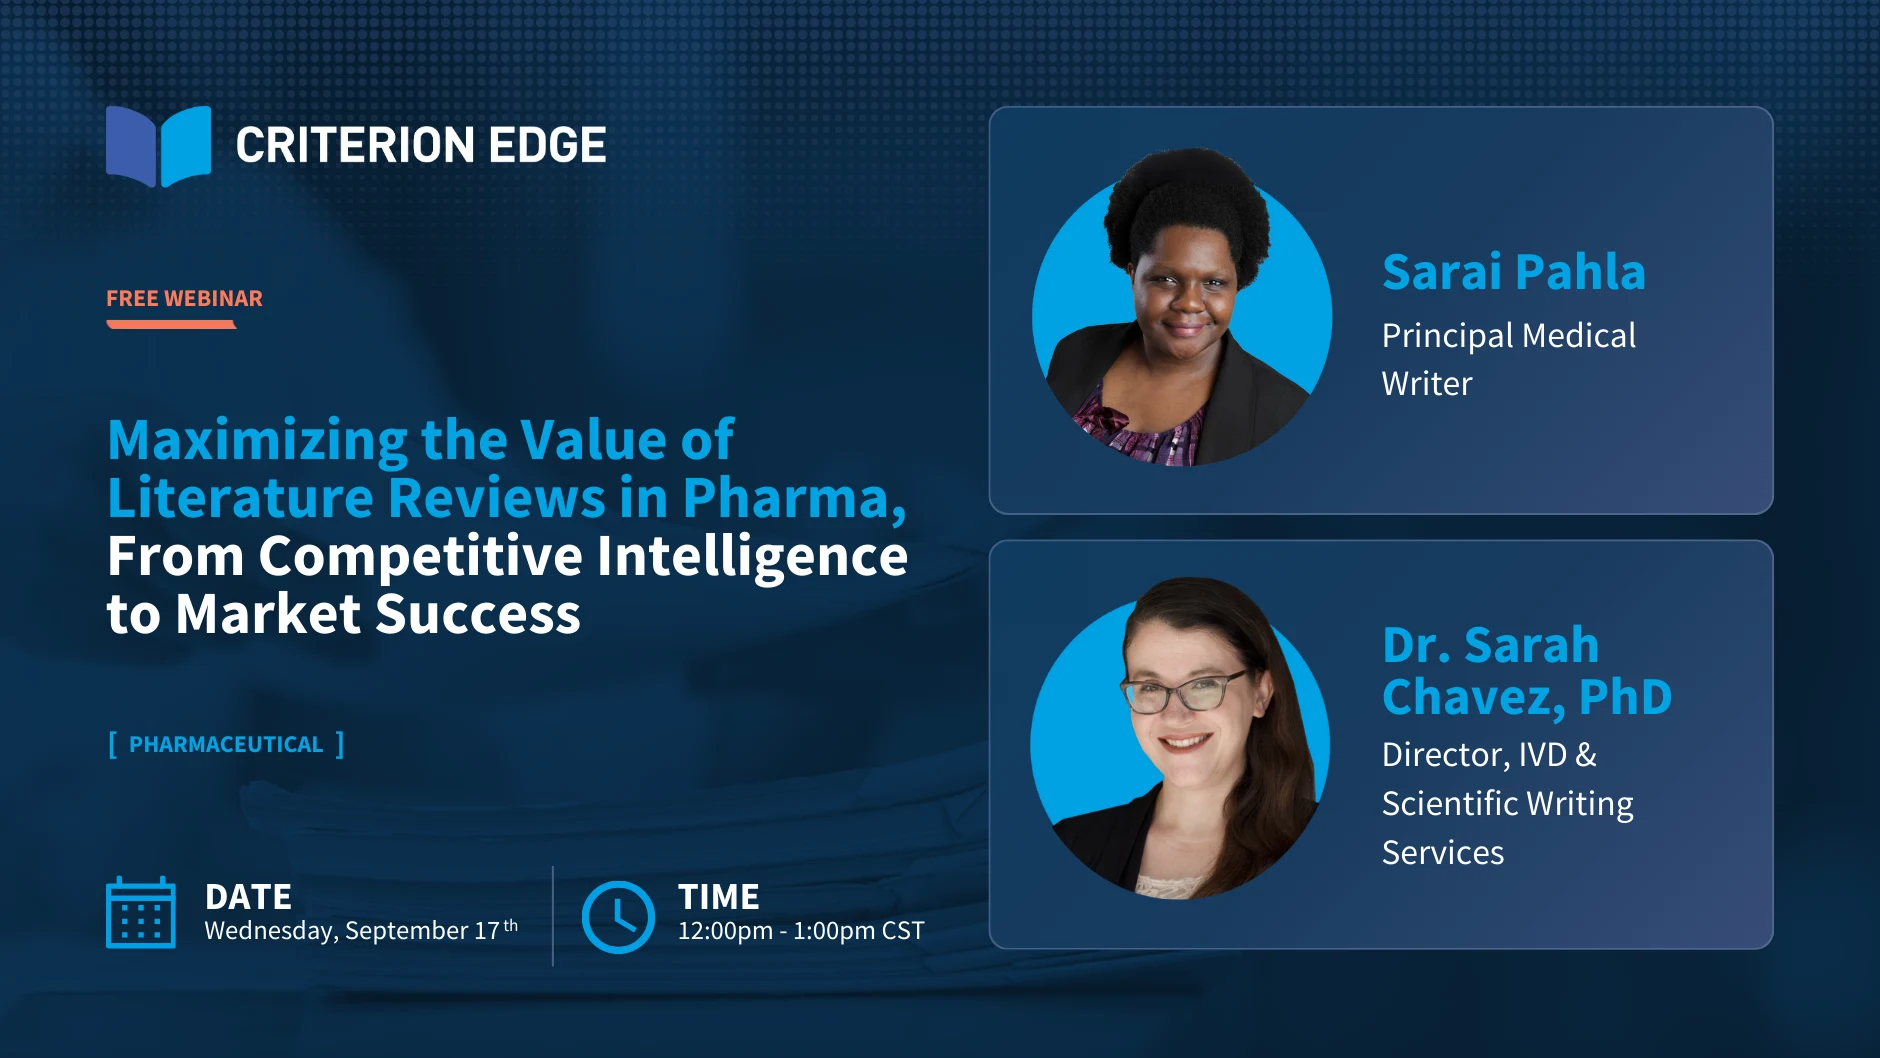
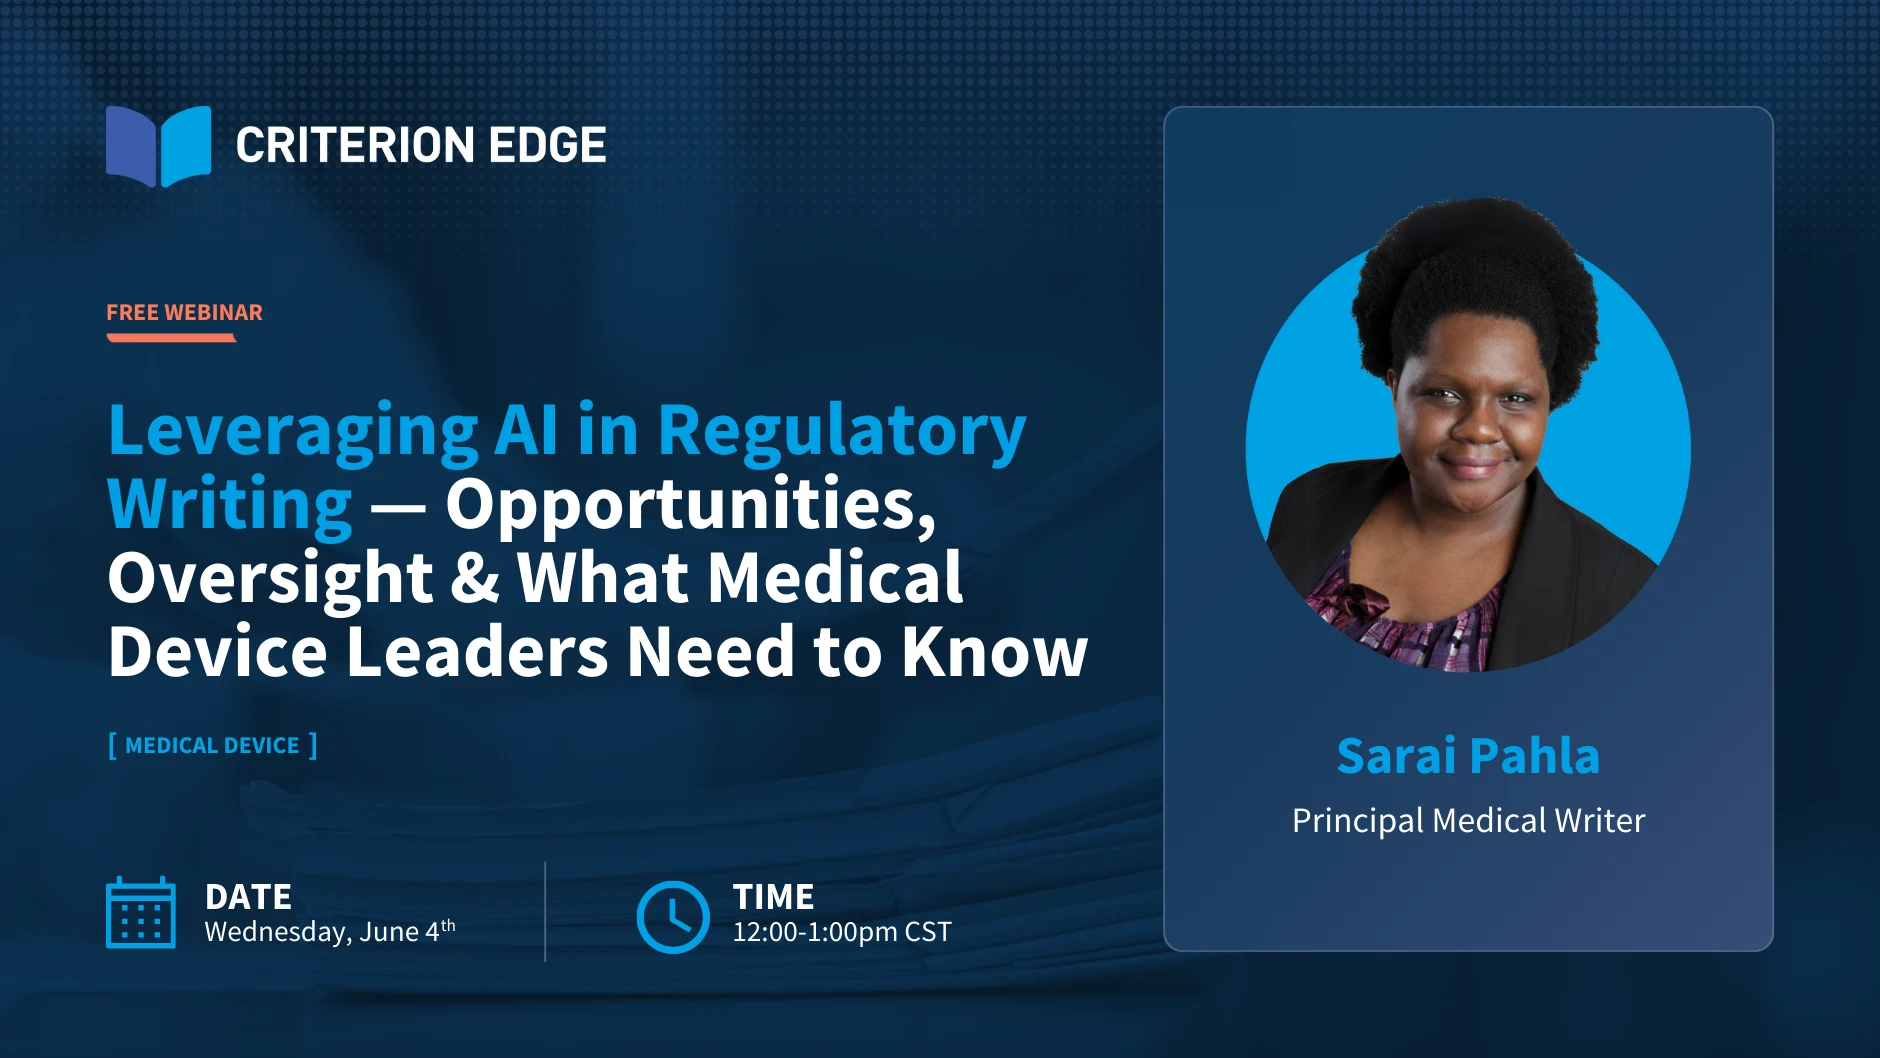
Pharmaceutical and medical device companies face growing pressure to deliver high-quality, compliant evidence on tight timelines - with limited internal resources. Criterion Edge was built to fill that gap with expert-led, 510(k) services.
Meeting FDA 510(k) Standards With Confidence From eSTAR compliance to substantial equivalence documentation and evolving FDA guidance, regulatory requirements demand expertise and vigilance. We navigate the pathway to clearance for you.
Late reports, misaligned expectations, or unclear deliverables waste valuable time. Our team is known for quality, clarity, and on-time delivery.
Internal 510(k) teams can be costly and hard to scale. Outsourcing with us helps you meet your evidence goals while staying within budget.
510(k) expertise is niche and hard to hire. Our experienced writers and reviewers are ready to support your projects right now.
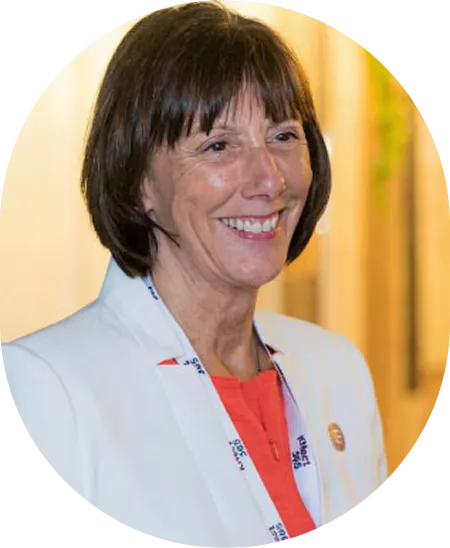
With over two decades in the pharmaceutical and medical device sectors, Laurie Mitchell saw firsthand how regulatory and clinical teams struggled with the complexities of 510(k) submissions. She launched Criterion Edge to deliver high-quality regulatory evidence and documentation that bridges the gap between technical requirements and submission-ready reports.
Under her leadership, Criterion Edge has supported top global brands across 510(k) submissions, predicate device analysis, and regulatory strategy—providing expert-written, fully documented deliverables trusted by teams around the world.
Let’s discuss your 510(k) needs and how Criterion Edge can support your strategic objectives.